
The affected products and recommendations for what to do with the devices below have not changed. This recall involves removing certain devices from where they are used or sold. The FDA has identified this recall as the most serious type. This device may cause serious injury or death if you continue to use it.
Affected Product
The FDA is aware that Draeger has issued a letter to affected customers recommending certain VentStar Resus Neo hoses be removed from where they are used or sold.
Affected devices:
|
Product Name |
Model |
UDI |
|---|---|---|
|
VentStar Resus Neo |
MP00310 |
What to Do
These devices are used in critical and emergency situations. Users should review and follow the recommendations below to inspect and remove affected product prior to use.
On September 3, Draeger sent all affected customers a letter recommending the following actions:
- Carefully inspect the hoses for damages prior to use according to the product’s Instructions for Use (IFU).
- Discard products that show cracks (Figure 1) or have a detached hose (Figure 2). You do not need to return them to Draeger.
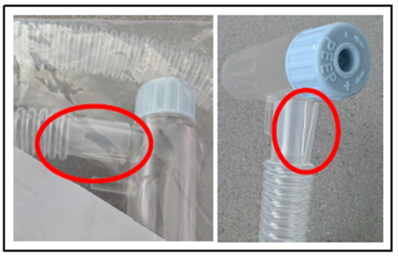
Figure 1. Cracked hose.
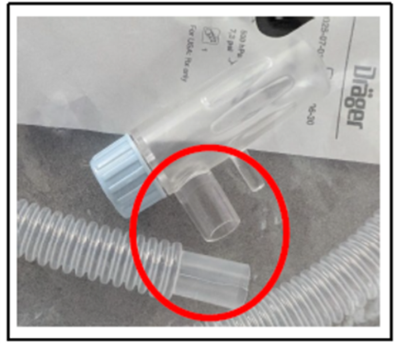
Figure 2. Detached hose.
Reason for Recall
Draeger stated that they are aware of cases in which obvious cracks in the hose of the VentStar Resus Neo were detected by users prior to use.
If a cracked hose is used, the hose may fall off or leak, resulting in restricted ventilation or delayed therapy which could lead to desaturation, hypoxia, or patient death.
As of September 3, Draeger has reported no serious injuries or deaths associated with this issue.
Device Use
The VentStar Resus Neo is a disposable breathing circuit used to convey breathing gas from a source to an infant (up to 22 lbs).
Contact Information
Customers in the U.S. with adverse reactions, quality problems, or questions about this issue should contact Draeger at 445-224-0698 or karina.howard@draeger.com.
Additional FDA Resources
Unique Device Identifier (UDI)
The unique device identifier (UDI) helps identify individual medical devices sold in the United States from distribution to use. The UDI allows for more accurate reporting, reviewing, and analyzing of adverse event reports so that devices can be identified more quickly, and as a result, problems potentially resolved more quickly.
- How do I recognize a UDI on a label?
- AccessGUDID database – Identify Your Medical Device
- Benefits of a UDI System
How do I report a problem?
Health care professionals and consumers may report adverse reactions or quality problems they experienced using these devices to MedWatch: The FDA Safety Information and Adverse Event Reporting Program.
Timeline of Communication Updates
| Date | Actions |
|---|---|
| 10/02/2026 | The FDA updated this communication to inform the public that this issue has been classified as a Class I Recall. |
| 09/03/2026 | The FDA issued an Early Alert communication to notify the public of a potentially high-risk device issue. |